
产品中心
- 细胞类
- 生化试剂
- ELISA检测
-
抗体蛋白
二抗生物素标记 过氧化物酶(HRP)标记 胶体金试剂 FITC荧光标记 RBITC荧光标记 二抗免疫血清 其它荧光标记二抗 藻红蛋白(PE)荧光标记 胶体金(Gold)标记 SAlexa Fluor荧光系列 碱性磷酸酶(AP)标记 别藻蓝蛋白(APC)荧光标记 其它标记 PE标记二抗 DyLight标记二抗 AU标记二抗 Biotin标记二抗 AMCA标记二抗 Texas Red标记二抗 TRITC标记二抗 HRP标记二抗 未标记二抗 Cy标记二抗 AbBox Fluor标记二抗内参抗体 小分子抗体抗体标记试剂盒细菌抗体蛋白病毒包装试剂杂交瘤融合筛选WB、IHC、ELISA相关试剂细胞培养试剂病原微生物抗原抗体假病毒抗体校准品其他抗原抗体标记的标签抗体病理级IHC抗体重组蛋白
- 细胞培养
- 实验耗材
- 仪器设备
- 生化试剂盒
- 小分子试剂
- 基质胶
-
斑马鱼产品
订货方式
订货电话:13564444959
订货时间:周一至周五
订货Q Q:79688691
订货邮件:79688691@qq.com
订货时间:周一至周五
订货Q Q:79688691
订货邮件:79688691@qq.com
产品简介
储存与保存
注意事项
 产品简介
-
产品名称Collagen I α1 (Cleaved Ala1218) Rabbit Polyclonal Antibody
-
别名Collagen alpha-1(I) chain (Alpha-1 type I collagen)
-
类别抗体产品
-
基因名称COL1A1
-
蛋白名称Collagen I α1 (Cleaved-Ala1218)
-
ClonalityPolyclonal
-
推荐应用WB,ELISA
-
稀释度WB 1:1000-2000, ELISA 1:5000-20000
-
反应种属Human,Mouse,Rat
-
浓度1 mg/ml
-
存储缓冲液Liquid in PBS containing 50% glycerol, 0.5% BSA and 0.02% New type preservative N.
-
Human Gene ID1277
-
Human Swissprot No.P02452
-
Mouse Gene ID12842
-
Mouse Swissprot No.P11087
-
Rat Gene ID29393
-
Rat Swissprot No.P02454
-
免疫原Synthesized peptide derived from human Collagen I α1 (Cleaved-Ala1218)
-
特异性This antibody detects endogenous levels of Human,Mouse,Rat Collagen I α1 (Cleaved-Ala1218, protein was cleaved amino acid sequence between 1218-1219 )
-
预测分子量135 160kDa
-
宿主Rabbit
-
同种型Rabbit,IgG
-
背景介绍This gene encodes the pro-alpha1 chains of type I collagen whose triple helix comprises two alpha1 chains and one alpha2 chain. Type I is a fibril-forming collagen found in most connective tissues and is abundant in bone, cornea, dermis and tendon. Mutations in this gene are associated with osteogenesis imperfecta types I-IV, Ehlers-Danlos syndrome type VIIA, Ehlers-Danlos syndrome Classical type, Caffey Disease and idiopathic osteoporosis. Reciprocal translocations between chromosomes 17 and 22, where this gene and the gene for platelet-derived growth factor beta are located, are associated with a particular type of skin tumor called dermatofibrosarcoma protuberans, resulting from unregulated expression of the growth factor. Two transcripts, resulting from the use of alternate polyadenylation signals, have been identified for this gene. [provided by R. Dalgleish, Feb 2008],
-
组织表达Forms the fibrils of tendon, ligaments and bones. In bones the fibrils are mineralized with calcium hydroxyapatite.
-
细胞定位Secreted, extracellular space, extracellular matrix
-
功能disease:A chromosomal aberration involving COL1A1 is a cause of dermatofibrosarcoma protuberans (DFSP) [MIM:607907]. Translocation t(17;22)(q22;q13) with PDGF. DFSP is an uncommon, locally aggressive, but rarely metastasizing tumor of the deep dermis and subcutaneous tissue. It typically occurs during early or middle adult life and is most frequently located on the trunk and proximal extremities.,disease:Defects in COL1A1 are a cause of Ehlers-Danlos syndrome type 1 (EDS1) [MIM:130000]; also known as Ehlers-Danlos syndrome gravis. EDS is a connective tissue disorder characterized by hyperextensible skin, atrophic cutaneous scars due to tissue fragility and joint hyperlaxity. EDS1 is the severe form of classic Ehlers-Danlos syndrome.,disease:Defects in COL1A1 are a cause of osteogenesis imperfecta type I (OI-I) [MIM:166200]. OI-I is a dominantly inherited serious newborn disease characterized by bone fragility, normal stature, little or no deformity, blue sclerae and hearing loss in 50% of families. Dentinogenesis imperfecta is rare and may distinguish a subset of OI type I (formation of dentine).,disease:Defects in COL1A1 are a cause of osteogenesis imperfecta type II (OI-II) [MIM:166210]; also known as osteogenesis imperfecta congenita. OI-II is lethal in the perinatal period and is charaterized by calvarial mineralization, beaded ribs, compressed femurs, marked long bone deformity and platyspondyly (congenital flattening of the vertebral bodies).,disease:Defects in COL1A1 are a cause of osteogenesis imperfecta type III (OI-III) [MIM:259420]; also called progressively deforming osteogenesis imperfecta with normal sclerae. OI-III is characterized by progressively deforming bones, usually with moderate deformity at birth, sclerae is variable in color, dentinogenesis imperfecta and hearing loss are common. The stature is very short.,disease:Defects in COL1A1 are a cause of osteogenesis imperfecta type IV (OI-IV) [MIM:166220]. OI-IV is charaterized by normal sclerae, moderate to mild deformity and variable short stature. Dentinogenesis imperfecta is common and hearing loss occurs in some patients.,disease:Defects in COL1A1 are the cause of Caffey disease [MIM:114000]; also known as infantile cortical hyperostosis. Caffey disease is characterized by an infantile episode of massive subperiosteal new bone formation that typically involves the diaphyses of the long bones, mandible, and clavicles. The involved bones may also appear inflamed, with painful swelling and systemic fever often accompanying the illness. The bone changes usually begin before 5 months of age and resolve before 2 years of age.,disease:Defects in COL1A1 are the cause of Ehlers-Danlos syndrome type 7A (EDS7A) [MIM:130060]; also known as autosomal dominant Ehlers-Danlos syndrome type VII. EDS is a connective tissue disorder characterized by hyperextensible skin, atrophic cutaneous scars due to tissue fragility and joint hyperlaxity. EDS7A is marked by bilateral congenital hip dislocation, hyperlaxity of the joints, and recurrent partial dislocations.,disease:Genetic variations in COL1A1 are associated with susceptibility to involutional osteoporosis [MIM:166710]; also known as senile osteoporosis or postmenopausal osteoporosis. Osteoporosis is characterized by reduced bone mineral density, disrutption of bone microarchitecture, and the alteration of the amount and variety of non-collagenous proteins in bone. Osteoporotic bones are more at risk of fracture.,function:Type I collagen is a member of group I collagen (fibrillar forming collagen).,online information:Collagen type I alpha-1 chain mutations,online information:Type-I collagen entry,PTM:O-linked glycan consists of a Glc-Gal disaccharide bound to the oxygen atom of a post-translationally added hydroxyl group.,PTM:Proline residues at the third position of the tripeptide repeating unit (G-X-Y) are hydroxylated in some or all of the chains.,similarity:Belongs to the fibrillar collagen family.,similarity:Contains 1 VWFC domain.,subunit:Trimers of one alpha 2(I) and two alpha 1(I) chains. Interacts with MRC2.,tissue specificity:Forms the fibrils of tendon, ligaments and bones. In bones the fibrils are mineralized with calcium hydroxyapatite.,
-
纯化The antibody was affinity-purified from rabbit serum by affinity-chromatography using specific immunogen.
储存与保存1.保存:-20℃,Avoid freeze/thaw cycles
2.有效期:1年
注意事项1. 本产品仅供科研使用。请勿用于医药、临床诊断或治疗,食品及化妆品等用途。请勿存放于普通住宅区。
2. 为了您的安全和健康,请穿好实验服并佩戴一次性手套和口罩操作。
3. 实验结果可由多种因素影响,相关处理只限于产品本身,不涉及其他赔偿。
备注:由于产品信息可能会有优化升级,请以实际收货标签信息为准。
产品推荐
-
 β-Tubulin Mouse Monoclonal Antibody,HRP Conjugated¥2200.00货号:IM70308
-
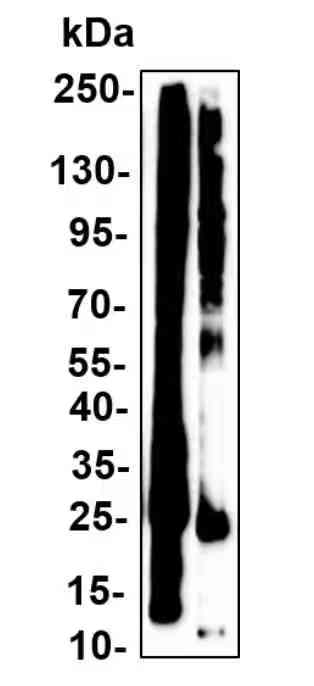 L-Lactyl Lysine (17E10) Rabbit Monoclonal Antibody¥1600.00货号:IM39531
-
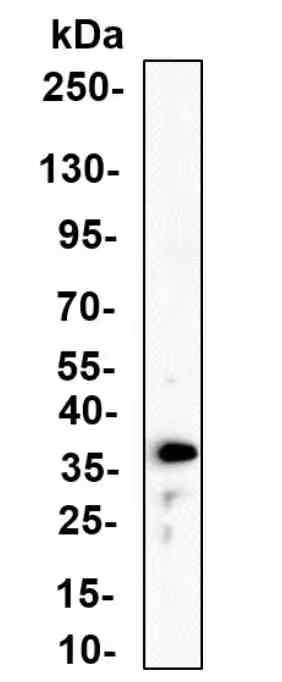 Annexin II (5W1) Rabbit Monoclonal Antibody¥1600.00货号:IM35141
-
 p300 (12P7) Rabbit Monoclonal Antibody¥1600.00货号:IM35111
-
 EHMT2 (1R17) Rabbit Monoclonal Antibody¥1600.00货号:IM35217