
产品中心
- 细胞类
- 生化试剂
- ELISA检测
-
抗体蛋白
二抗生物素标记 过氧化物酶(HRP)标记 胶体金试剂 FITC荧光标记 RBITC荧光标记 二抗免疫血清 其它荧光标记二抗 藻红蛋白(PE)荧光标记 胶体金(Gold)标记 SAlexa Fluor荧光系列 碱性磷酸酶(AP)标记 别藻蓝蛋白(APC)荧光标记 其它标记 PE标记二抗 DyLight标记二抗 AU标记二抗 Biotin标记二抗 AMCA标记二抗 Texas Red标记二抗 TRITC标记二抗 HRP标记二抗 未标记二抗 Cy标记二抗 AbBox Fluor标记二抗内参抗体 小分子抗体抗体标记试剂盒细菌抗体蛋白病毒包装试剂杂交瘤融合筛选WB、IHC、ELISA相关试剂细胞培养试剂病原微生物抗原抗体假病毒抗体校准品其他抗原抗体标记的标签抗体病理级IHC抗体重组蛋白
- 细胞培养
- 实验耗材
- 仪器设备
- 生化试剂盒
- 小分子试剂
- 基质胶
-
斑马鱼产品
订货方式
订货电话:13564444959
订货时间:周一至周五
订货Q Q:79688691
订货邮件:79688691@qq.com
订货时间:周一至周五
订货Q Q:79688691
订货邮件:79688691@qq.com
产品简介
储存与保存
注意事项
 产品简介
-
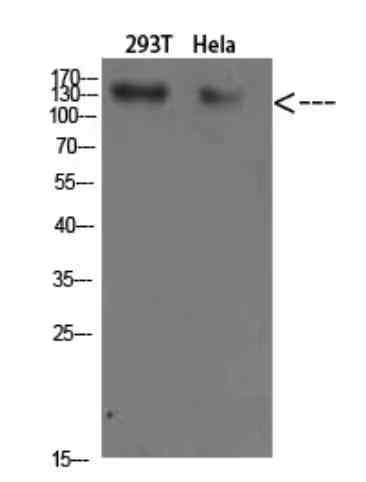
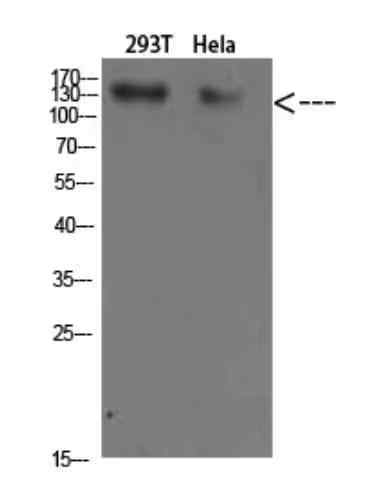
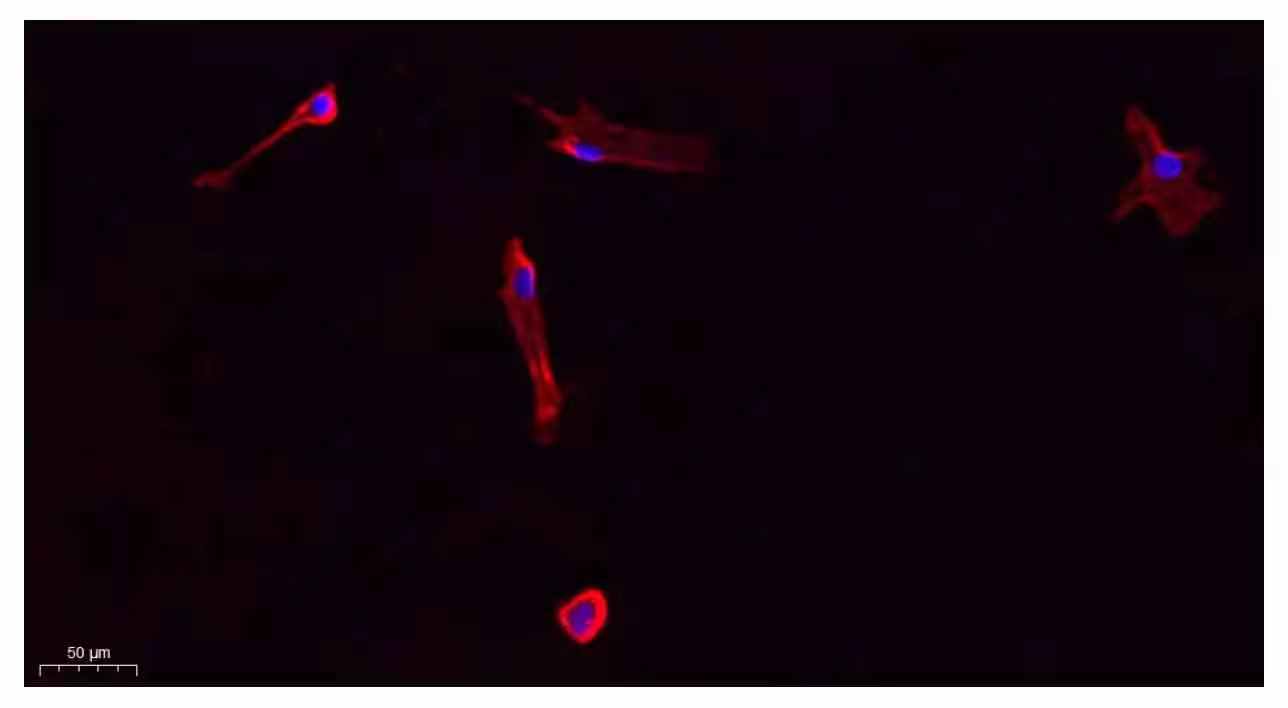
产品名称Collagen I Rabbit Polyclonal Antibody
-
别名collagen, type I, alpha 2
-
类别抗体产品
-
基因名称Collagen I
-
ClonalityPolyclonal
-
推荐应用WB IF-P,IF-F,IF-ICC,ELISA
-
稀释度WB 1:500-2000, IF-P/IF-F/IF-ICC 1:100-300, ELISA 1:5000-20000, Not yet tested in other applications.
-
反应种属Human,Mouse,Rat
-
浓度1 mg/ml
-
存储缓冲液Liquid in PBS containing 50% glycerol, 0.5% BSA and 0.02% New type preservative N.
-
Human Gene ID1278
-
Human Swissprot No.P08123/P02452
-
免疫原Synthesized peptide derived from Collagen I . at AA range: 1200-1217
-
特异性Collagen I Polyclonal Antibody detects endogenous levels of Collagen I
-
预测分子量135kDa
-
宿主Rabbit
-
同种型Rabbit,IgG
-
背景介绍This gene encodes the pro-alpha2 chain of type I collagen whose triple helix comprises two alpha1 chains and one alpha2 chain. Type I is a fibril-forming collagen found in most connective tissues and is abundant in bone, cornea, dermis and tendon. Mutations in this gene are associated with osteogenesis imperfecta types I-IV, Ehlers-Danlos syndrome type VIIB, recessive Ehlers-Danlos syndrome Classical type, idiopathic osteoporosis, and atypical Marfan syndrome. Symptoms associated with mutations in this gene, however, tend to be less severe than mutations in the gene for the alpha1 chain of type I collagen (COL1A1) reflecting the different role of alpha2 chains in matrix integrity. Three transcripts, resulting from the use of alternate polyadenylation signals, have been identified for this gene. [provided by R. Dalgleish, Feb 2008],
-
组织表达Forms the fibrils of tendon, ligaments and bones. In bones the fibrils are mineralized with calcium hydroxyapatite.
-
细胞定位Secreted, extracellular space, extracellular matrix .
-
信号通路Focal adhesion;ECM-receptor interaction;
-
功能disease:A chromosomal rearrangement involving COL1A2 may be a cause of lipoblastomas, which are benign tumors resulting from transformation of adipocytes, usually diagnosed in children. Translocation t(7;8)(p22;q13) with PLAG1.,disease:Defects in COL1A2 are a cause of osteogenesis imperfecta type I (OI-I) [MIM:166200]. OI-I is a dominantly inherited serious newborn disease characterized by bone fragility, normal stature, little or no deformity, blue sclerae and hearing loss in 50% of families. Dentinogenesis imperfecta is rare and may distinguish a subset of OI type I (formation of dentine).,disease:Defects in COL1A2 are a cause of osteogenesis imperfecta type II (OI-II) [MIM:166210]; also known as osteogenesis imperfecta congenita (OIC) or lethal perinatal. OI-II is a serious newborn disease that diffusely affects bone. Infants are born with multiple fractures, which lead to shortening of the extremities. The skull is soft, and resembles a "bag of bones" when palpated, the sclera are abnormally thin and may appear blue, and some infants also have a hearing loss. Infants born alive often die suddenly during the first few days or weeks of life, but a few survive as deformed dwarfs. Mental development is normal unless head trauma with CNS injury occurs. There is no effective treatment.,disease:Defects in COL1A2 are a cause of osteogenesis imperfecta type III (OI-III) [MIM:259420]. OI-III usually presents with moderate deformity at birth, progressively deforming bones, and sclerae variable in color. Dentinogenesis imperfecta and hearing loss are common. Stature is very short.,disease:Defects in COL1A2 are a cause of osteogenesis imperfecta type IV (OI-IV) [MIM:166220]; also known as osteogenesis imperfecta with normal sclerae. OI-IV presents with moderate to mild deformity and variable short stature. Dentinogenesis imperfecta is common and hearing loss occurs in some.,disease:Defects in COL1A2 are the cause of cardiac valvular form of autosomal recessive Ehlers-Danlos syndrome (cardiac valvular EDS) [MIM:225320]; also known as arthrochalasis type Ehlers-Danlos syndrome. In addition to joint laxity, skin hyperextensibility and friability, and abnormal scar formation, individuals with this form of EDS appear to be at increased risk for cardiac valvular dysfunction.,disease:Defects in COL1A2 are the cause of Ehlers-Danlos syndrome type 7B (EDS7B) [MIM:130060]. EDS is a connective tissue disorder characterized by hyperextensible skin, atrophic cutaneous scars due to tissue fragility and joint hyperlaxity. EDS7B is marked by bilateral congenital hip dislocation, hyperlaxity of the joints, and recurrent partial dislocations.,function:Type I collagen is a member of group I collagen (fibrillar forming collagen).,online information:Collagen type I alpha-2 chain mutations,PTM:Prolines at the third position of the tripeptide repeating unit (G-X-Y) are hydroxylated in some or all of the chains.,similarity:Belongs to the fibrillar collagen family.,subunit:Trimers of one alpha 2(I) and two alpha 1(I) chains.,tissue specificity:Forms the fibrils of tendon, ligaments and bones. In bones the fibrils are mineralized with calcium hydroxyapatite.,
-
纯化The antibody was affinity-purified from rabbit antiserum by affinity-chromatography using epitope-specific immunogen.
储存与保存1.保存:-20℃,Avoid freeze/thaw cycles
2.有效期:1年
注意事项1. 本产品仅供科研使用。请勿用于医药、临床诊断或治疗,食品及化妆品等用途。请勿存放于普通住宅区。
2. 为了您的安全和健康,请穿好实验服并佩戴一次性手套和口罩操作。
3. 实验结果可由多种因素影响,相关处理只限于产品本身,不涉及其他赔偿。
备注:由于产品信息可能会有优化升级,请以实际收货标签信息为准。
产品推荐
-
 Annexin II (5W1) Rabbit Monoclonal Antibody¥1600.00货号:IM35141
-
 p300 (12P7) Rabbit Monoclonal Antibody¥1600.00货号:IM35111
-
 EHMT2 (1R17) Rabbit Monoclonal Antibody¥1600.00货号:IM35217
-
 eNOS (Phospho Ser1177) (17W2) Rabbit Monoclonal Antibody¥1600.00货号:IM34470
-
 Glucagon (19V15) Rabbit Monoclonal Antibody¥1600.00货号:IM33382